RNA sequencing: which genes changed, and what that means for your hypothesis.
We measure the activity of every gene in your samples, compare your groups and connect the changed genes to biological pathways.
Counting RNA copies shows how active each gene is.
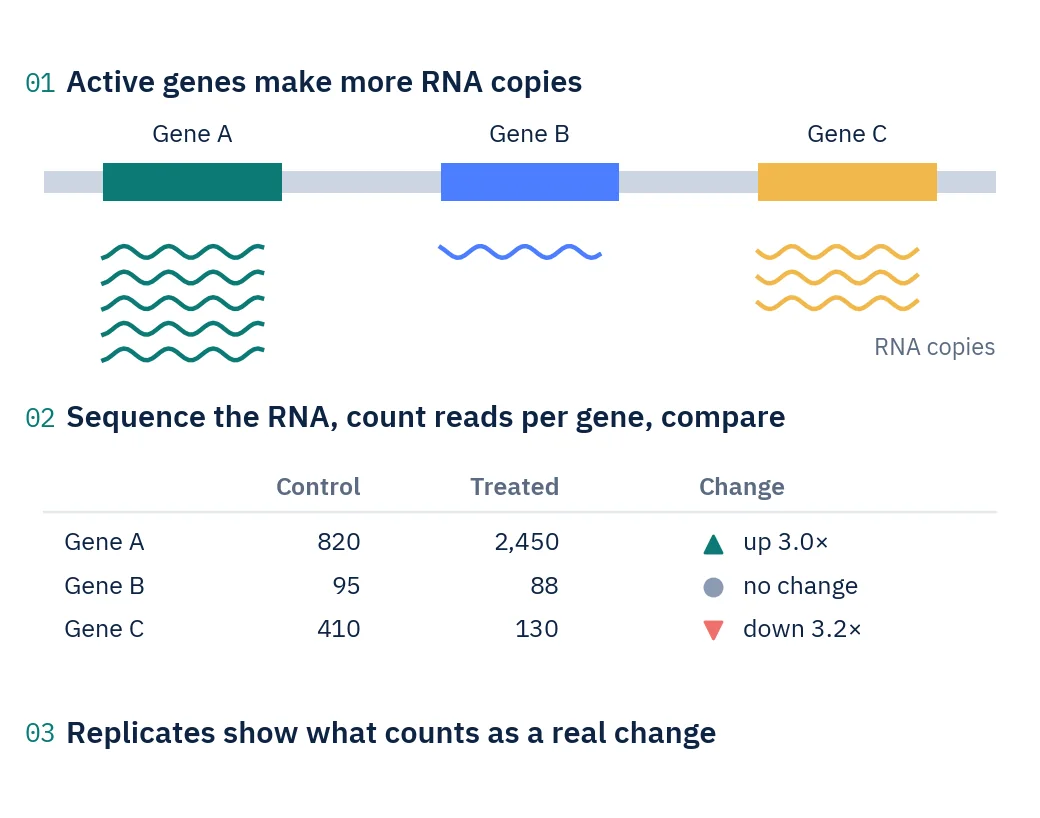
A gene that is switched on is copied into messenger RNA, and a busy gene makes many copies. RNA sequencing reads millions of these molecules from each sample and counts how many come from each gene. That count measures gene activity at the moment the sample was taken.
Most RNA in a cell is ribosomal RNA, which says little about gene activity. Before sequencing, the library is focused on the informative part: messenger RNA is captured by its poly(A) tail, or ribosomal RNA is removed so that non-coding RNAs and bacterial transcripts stay in the data.
Samples receive different numbers of reads, so counts are normalised before groups are compared. A gene counts as changed when the difference between groups is clearly larger than the natural variation between biological replicates, after correcting for testing about 20,000 genes at once.
Key terms
- Transcriptome
- All the RNA molecules in a sample at one moment.
- mRNA
- The RNA copy of a gene that the cell uses to make a protein.
- Biological replicate
- An independent sample from the same group, such as another animal or culture. Replicates measure natural variation.
- Normalisation
- Scaling counts so that samples sequenced to different depths can be compared fairly.
- Log2 fold change
- How much a gene changed on a doubling scale: +1 means twice as much, −1 means half.
- Adjusted p-value
- A p-value corrected for testing thousands of genes at once, which keeps false discoveries in check.
Bring the question. We design the experiment around it.
- Which genes respond to the treatment, and how strongly?
- Which biological pathways and processes are affected?
- Do my samples group by condition, or is an outlier driving the result?
- Which of several treatments differ from the control?
Where it is used
- Treatment and disease studiesGene activity in treated versus control, or diseased versus healthy samples.
- Mechanism and pathway researchLinking expression changes to biological processes and signalling pathways.
- Biomarker discoveryGenes whose activity separates groups or tracks a response.
- Plants and cropsStress responses and trait research, including species beyond the usual model organisms.
- Host and virus togetherHost gene activity and viral transcripts measured in the same infected samples.
- Organisms without a genomeA transcript catalogue assembled directly from the reads.
Four steps from RNA to results.
- Step 1
Plan the comparison
We agree on groups, replicates and the comparisons that answer your question. Three or more biological replicates per group is the usual minimum.
- Step 2
Check the RNA
RNA amount and integrity are checked before library preparation, so problems show up early.
- Step 3
Build libraries and sequence
mRNA is captured or ribosomal RNA removed, and the libraries are sequenced as 150-letter read pairs.
- Step 4
Analyse and report
Reads are placed on the genome and counted per gene, then groups are compared and pathways tested, with figures and a Methods section.
What the analysis shows you.
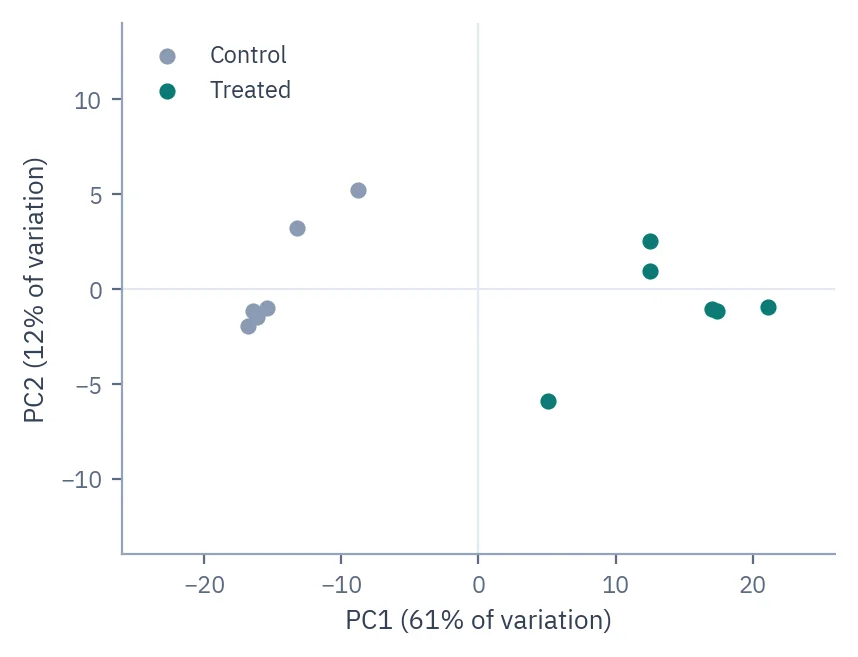
Do the samples group?
Each dot is one sample. Replicates sit together and the treatment separates the groups along the first axis, a sign of consistent data.
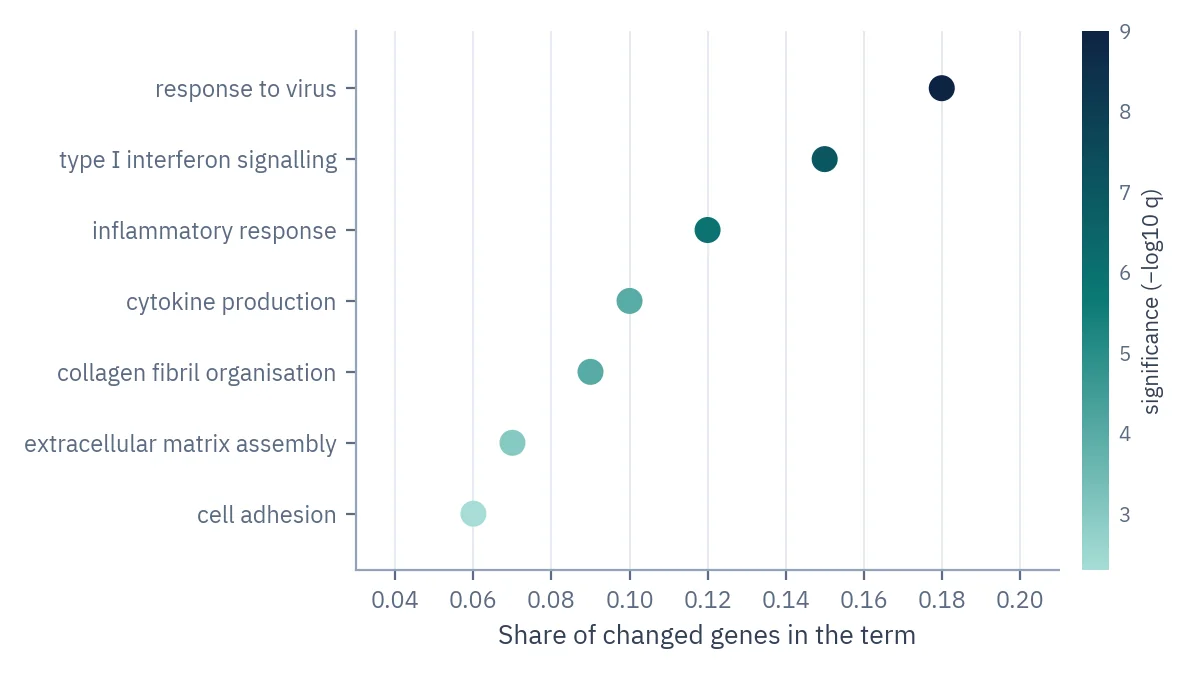
Pathways behind the change
Biological processes that appear more often among the changed genes than chance would allow. Darker dots mean stronger evidence.
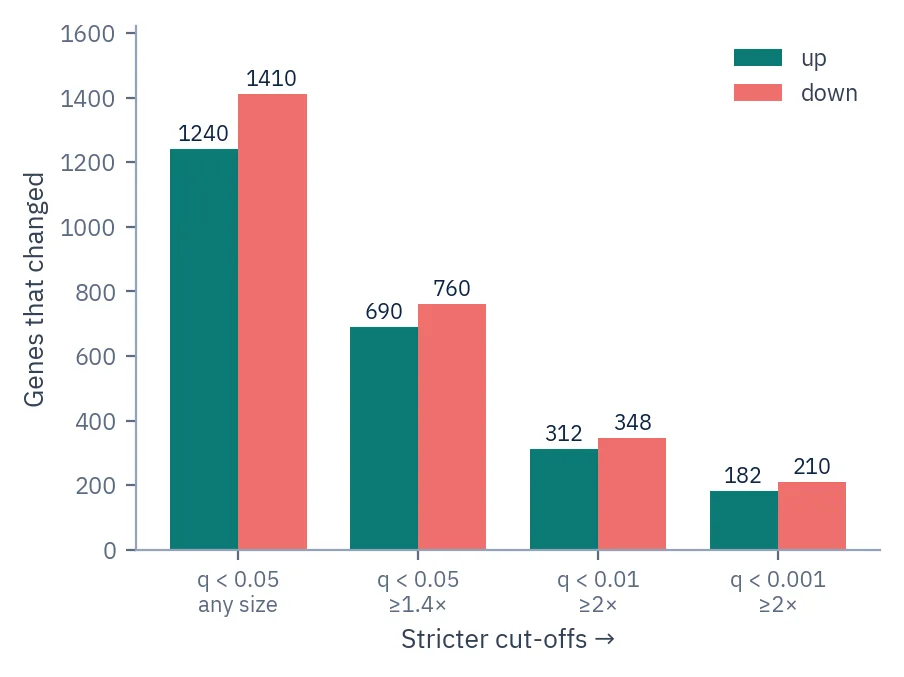
How robust is the gene list?
The number of changed genes at stricter and stricter cut-offs, so you see how much of the result holds up.
Included as standard every project
- Read quality controlAdapters and low-quality bases are removed, and every sample is checked.
- Alignment and gene countsReads are placed on the reference genome and counted per gene.
- Sample overviewPCA and sample-distance plots show whether replicates agree.
- Differential expressionChanged genes with fold change and adjusted p-value, at several cut-offs.
- Pathway enrichmentGO and KEGG terms over-represented among the changed genes.
Added for your question custom
- Several groupsSeparate results for every comparison in multi-group designs.
- Non-model organismsSpecies beyond human, mouse and rat, with the genome and annotation set up for your project.
- De novo transcriptomeA transcript catalogue with a completeness check and gene function annotation, when no genome exists.
- Host and virus readsViral transcripts quantified alongside host genes in infected samples.
What goes in, and how we run it.
- Libraries
- Poly(A) mRNA · rRNA-depleted total RNA · small RNA
- Sequencing
- Illumina, 150 bp read pairs
- Depth
- Set per project from the question, the library type and the genome
- Replicates
- Three or more biological replicates per group
- Samples
- Cells, tissue, blood, plants and microbes · degraded and FFPE RNA through total RNA protocols
- Main tools
- STAR, Salmon, DESeq2, clusterProfiler
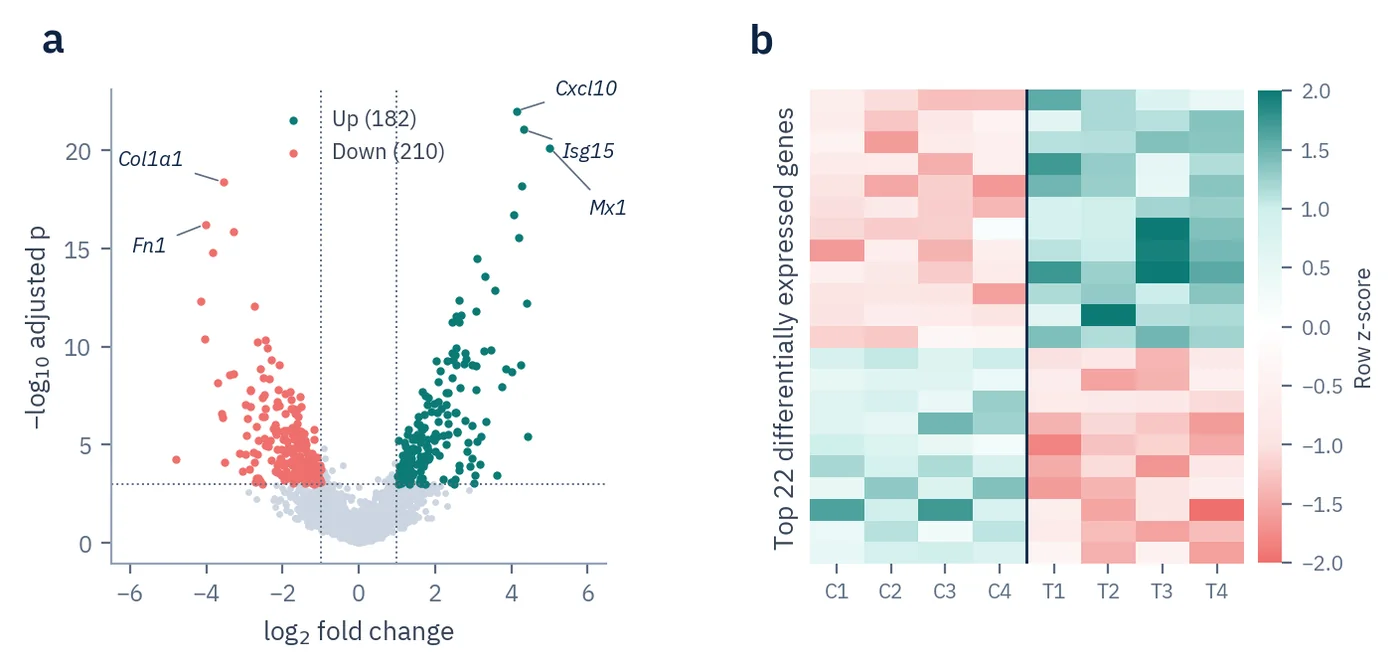
Figures, interpretation and Methods, ready for the manuscript.
- ReportInterpretation written against your hypothesis
- FiguresPublication-ready figures for the manuscript
- MethodsMaterials & Methods text for the paper
- Tables and dataAll results as tables, plus the processed data files
mRNA or total RNA?
Bring the question to the call and the scientist recommends one, with the reasoning for your samples.
| Aspect | mRNA sequencing | Total RNA sequencing |
|---|---|---|
| Captures | Messenger RNA with a poly(A) tail, mostly protein-coding genes | All RNA except ribosomal RNA, including non-coding RNA |
| Organisms | Animals, plants and fungi | Also bacteria, whose mRNA has no poly(A) tail |
| RNA quality | Works best with intact RNA | Copes with degraded and FFPE RNA |
| Reads needed | Fewer for gene-level answers | More for the same coverage of mRNA |
| Best when | Clean samples and questions about protein-coding genes | Degraded samples, non-coding RNA or bacteria |
Straight answers before you email.
Related services
How many biological replicates do I need?
Three per group is the minimum. With fewer than three there is no way to measure how much your samples naturally differ, and that is what the test compares against. Beyond three, the number depends on two things: how much your own samples vary from each other, and how small a change you need to detect. Neither can be guessed from the sample type, so we look at your pilot data or your earlier work. If your question needs more samples than your budget allows, we say so before you order, not after the data comes back.
How many reads per sample do I need?
It depends on what you want to measure, not on which organism you work with. Counting active genes needs fewer reads than finding rare transcripts, splice isoforms or new non-coding RNA. Libraries made by removing ribosomal RNA also use up some reads on material that the other method would have thrown away. We set the number from your question, your library type and your genome, and we write it into the quote so it does not go up later.
Can I send samples in several shipments, or add some later?
Yes, but treat it as part of the design, not just as shipping. Samples sequenced months apart can carry a technical difference, and it has to be checked for and corrected. If you already know the samples will arrive in waves, we plan the batches so each one holds all of your groups. If it comes up in the middle of a project, we tell you how much it weakens the result before you decide.
My effect is probably small. Will RNA sequencing see it?
That depends on how much your samples differ from each other and on how big the change is. It is not about the sequencing. The same number of samples can be plenty for one experiment and not enough for another, so there is no number anyone can give you without seeing your setup. At the design review we go through what you have and what you expect to find, and we tell you whether that combination can find it. We would rather say it before you spend than explain it afterwards.
Can you handle a time course, or more than two groups?
Yes. We fit multi-group and time-course studies as one model instead of running many separate two-group tests. That usually has more power to find real changes, and it handles multiple testing properly. You get a result for every comparison you asked for, and we agree that list before the analysis starts, so nobody is choosing comparisons after seeing the data.
What RNA quality do you need? Is there a RIN cut-off?
There is no fixed number that everyone agrees on. Anyone who gives you one is quoting their own kit, not a standard. What matters is whether enough of your RNA is still long enough to build a library from. We check that before library prep starts, and we tell you what your numbers mean for your plan.
How much RNA do I need to send?
There is no single number. It depends on the library type and on the state of your RNA, and there are kits made for very small amounts. We give you a figure for your samples rather than a general one. If you are close to the limit, send what you have and we will tell you after QC whether it works.
My samples are blood. Won’t globin take over the result?
It will, if nothing is done about it. In whole blood, globin can take up a large share of the reads and leave too few for the genes you care about. There is a step that removes it, in the same way ribosomal RNA is removed. Tell us the sample is blood and which tube you collected it in, and we choose the library for that.
My organism is a bacterium, so there is no poly(A) tail.
Then the poly(A) method cannot work, and we use total RNA with the ribosomal RNA removed. Bacterial ribosomal RNA needs different probes from the ones used for animals and plants, so the species matters. Tell us what the organism is and we will say whether a standard kit covers it or whether it needs something built for it.
Can you sequence host and pathogen from the same sample?
Yes, and it is usually better than separating them. We match the reads to both references and count them separately, so you see the host response and the pathogen activity from one library. The pathogen is often only a small part of the material, so we allow for that when we set the depth.
Does DNA contamination in my RNA matter?
Yes. It uses up reads, and it can make some genes look more active than they are. The normal answer is to treat the sample with DNase and then check that it worked, instead of assuming. We check this during QC, so a sample with leftover DNA gets dealt with before anything is built from it.
Can you extract the RNA, or do I have to send RNA?
Either works. Sending tissue removes one of the most common reasons RNA sequencing projects fail, which is RNA breaking down before it reaches the tube. If you prefer to extract it yourself, tell us the method you used. Column, TRIzol and bead kits keep different things, and that changes what the data can show.
mRNA or total RNA sequencing?
Use mRNA when your RNA is intact and you care about protein-coding genes. Use total RNA when you also want non-coding RNA, when your organism has no poly(A) tail, or when the RNA is damaged. Total RNA needs more reads to reach the same coverage of mRNA, because it keeps material that the other method removes. We choose from your samples and your question, not from a default setting.
Paired-end or single-end? What read length?
Counting how active genes are rarely needs long reads or both ends. Questions about splice isoforms, fusions, or building transcripts from scratch do need them, because those questions are about which pieces join together. We pick from your question, and you will know which case you are in before anything is ordered.
Do you use strand-specific libraries?
Yes, and for most projects it should be the normal choice. Without it you cannot tell a gene’s own RNA apart from an overlapping RNA read in the other direction. Wherever two genes overlap that way, the counts for both are wrong. If there is a reason not to use it on a project, we will tell you what it is.
Can you do small RNA or miRNA?
Yes, as a separate library. Small RNAs are lost both by the poly(A) method and by normal ribosomal RNA removal, so they need their own preparation and their own analysis. If you want mRNA and small RNA from the same samples, say so while we are planning, because it changes how much material you need to send.
What do I actually get back?
Cleaned reads, a count for every gene, a plot that shows whether your replicates agree, the list of changed genes at several cut-offs, and the biological pathways those genes belong to. You also get the figures and a Methods section written for a journal, plus any other analysis your project needs. The analysis is included in the project rather than quoted on top, and there is no limit on it. If a result raises a new question, we run it.
How do you decide a gene has changed?
We compare the difference between your groups with how much your replicates differ from each other. Then we correct the p-values, because thousands of genes are tested at once and some will look significant by chance. You get the result at several cut-offs instead of one, so you can see how much of the gene list survives a stricter test. Reviewers often ask for exactly that.
My species has no reference genome. Can you still do RNA sequencing?
Yes. We build a catalogue of transcripts from your reads and count against that. You also get a check of how complete that catalogue is, so you know how much of the expected gene set was found, and we add likely gene functions so the list means something. It usually needs more reads than mapping to a genome, and we plan for that.
Can you compare my data with a published dataset?
Yes, but there is something you should know first. Data made in another lab with another protocol carries a technical difference that is mixed up with the comparison you want to make. We can correct for part of it. We will tell you what the comparison can and cannot show, instead of handing you a figure that claims more than it should.
Is the analysis reproducible? Can someone else run it again?
Yes. The tool versions and settings go into the Methods section, and you get the input files as well as the results. If a reviewer or a collaborator wants to check the work, nothing in it only runs on our computers.
A reviewer asks for another analysis six months later. What then?
You ask, and we run it. Reviewer re-analysis is covered by the project guarantee and costs you nothing. If you asked us to keep your data, we hold it for up to five years, so the request does not start with you uploading a terabyte again. This covers new figures, different cut-offs, and comparisons you did not ask for the first time.
Start the project that ends in a paper.
One sentence will do. What the samples are and what you need to know. A scientist handles the rest.
- [email protected] Projects, samples and invoices
Start without committing to anything
- One sentence to start
- A short call to get the details
- A design, with the reasoning behind it
- Free shipping worldwide
- Free pre-analysis if you qualify
- Nothing starts until you agree